This guide shows how to register a single monomer from its SMILES, check the drawn structure, set its cap groups, and later correct it. It uses diethylglycine (Deg), an achiral dialkyl glycine used like Aib to stabilise helices, as the worked example, then an N-methylated norleucine whose cap group was left wrong to show what an edit looks like.
Prerequisites: the Manager or Administrator role in Biotoolkit Monomers. You need the monomer's structure as SMILES, CXSMILES or a molfile, and you need to know where it attaches to its neighbours.
Only your organisation's private monomers can be created, edited or deleted. A public monomer can be opened to read its definition and structure, but nothing in it can be changed. See Public Monomers.
Step 1: Write the SMILES with numbered attachment points
An attachment point is written as [*:n], where n is the R-group number. For a peptide backbone monomer, the convention is:
- R1 on the amine nitrogen (the N-terminal side),
- R2 on the carbonyl carbon (the C-terminal side),
- R3 on the side chain, if the monomer can branch.
Diethylglycine is glycine with two ethyl groups on the alpha carbon:
CCC(CC)(N[*:1])C([*:2])=O
The two [*:n] atoms stand for the bonds the monomer will make inside a chain, so the SMILES describes the residue, not the free amino acid. Write the structure with its full stereochemistry ([C@H], [C@@H], /, \) when it has any: the service compares structures on their canonical form, and a missing stereocentre makes a distinct monomer look like a duplicate of another one. Diethylglycine has no stereocentre, so there is nothing to add here.
See Attachment Points and CAP Groups for the other accepted notations and for CXSMILES. A centre that is racemic or whose configuration is not yet known is declared with a CXSMILES stereo group, |&1:n| or |o1:n|; see How to Resolve the Stereochemistry of a Monomer.
Step 2: Open the form
On the Monomers page, click New monomer. You can also start from the New monomer quick action of the Home page. The New monomer dialog opens with the definition fields on the left and the Structure panel on the right. The Definition tab is the only one on a new monomer; an existing monomer also has an Audit tab, and a Stereo tab when its structure carries stereochemistry to resolve.
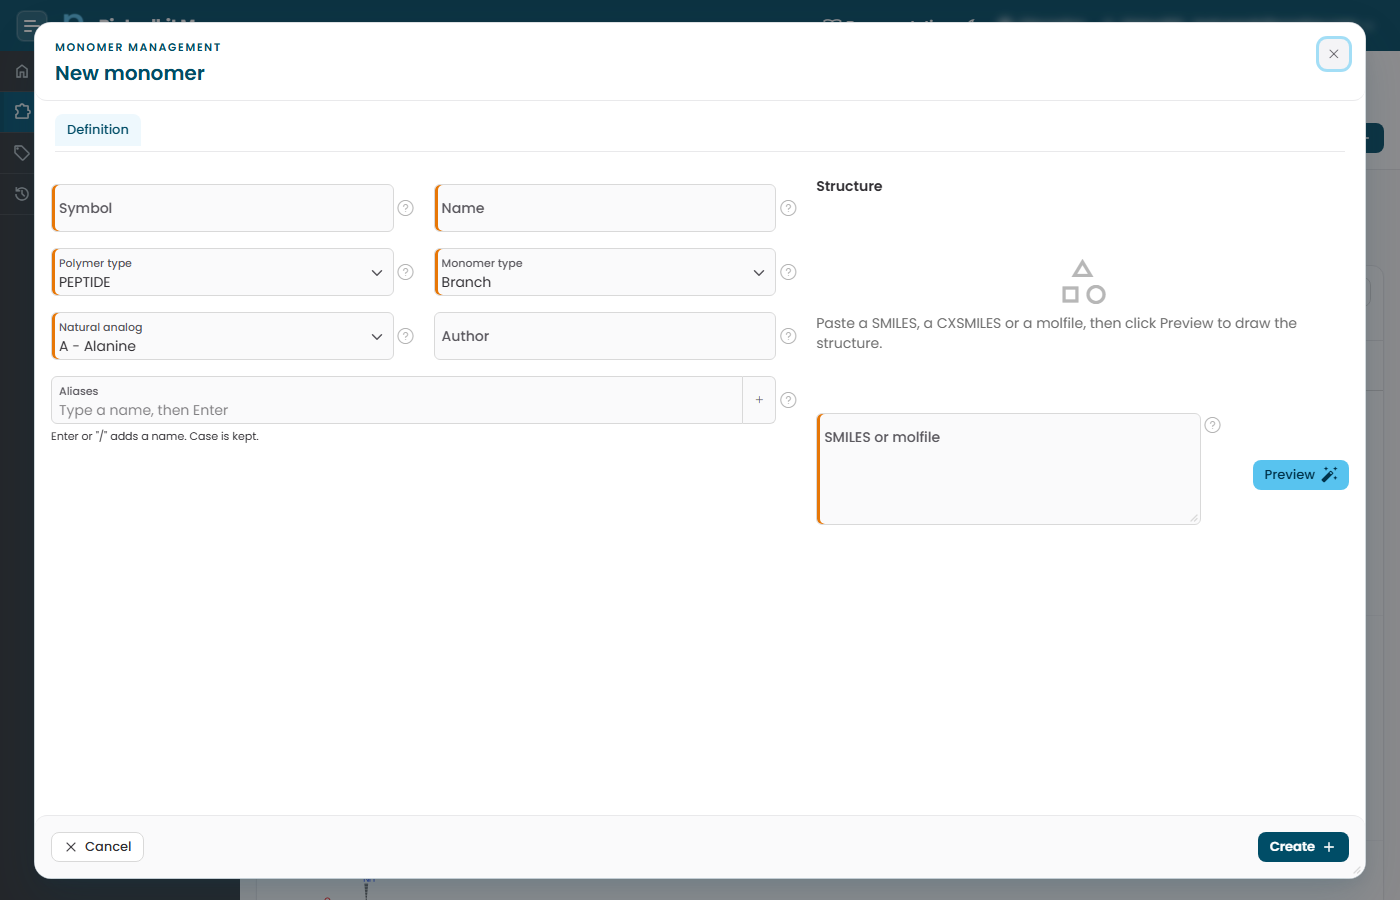
Step 3: Fill in the definition
| Field | Deg example | Notes |
|---|---|---|
| Symbol | Deg |
Required. Unique within your organisation for the chosen polymer type, and not already an alias. Cannot be changed once created. Case matters: Deg and DEG are two symbols. |
| Name | Diethylglycine |
Required. Unique within your organisation for the polymer type: a name another of your monomers already has is flagged under the field as soon as you leave it. |
| Polymer type | PEPTIDE |
Required, preselected. Cannot be changed once created. Selecting it filters the two lists below. |
| Monomer type | Backbone |
Required. Preselected on Backbone, a residue that sits in the chain; choose Branch for one that hangs off a side chain. |
| Natural analog | X - Unknown/Any |
Required. The natural residue this monomer stands in for. X - Unknown/Any is first in the list and preselected; a dialkyl glycine replaces no single natural residue, so keep it. |
| Author | your name | Optional free text. |
| Aliases | Et2Gly |
Optional. Other names the monomer answers to in HELM. Type a name, then press Enter, type / or click +. Each name is shown as a chip marked new and saved right after the monomer is created. See How to Manage Monomer Aliases. |
| SMILES or molfile | CCC(CC)(N[*:1])C([*:2])=O |
In the Structure panel. SMILES or CXSMILES with [*:n] attachment points, or a molfile. |
The meaning of each list, and which combinations exist, are described in Monomer Types, Polymer Types and Natural Analogs.
Step 4: Preview the structure
Click Preview, under the SMILES or molfile field. The application draws the molecule in the Structure panel with each attachment point labelled R1, R2, and so on, and an R groups section appears under the definition fields with one cap group selector per attachment point, two per row. The buttons above the drawing zoom it, and the last one opens it full size in the structure viewer.
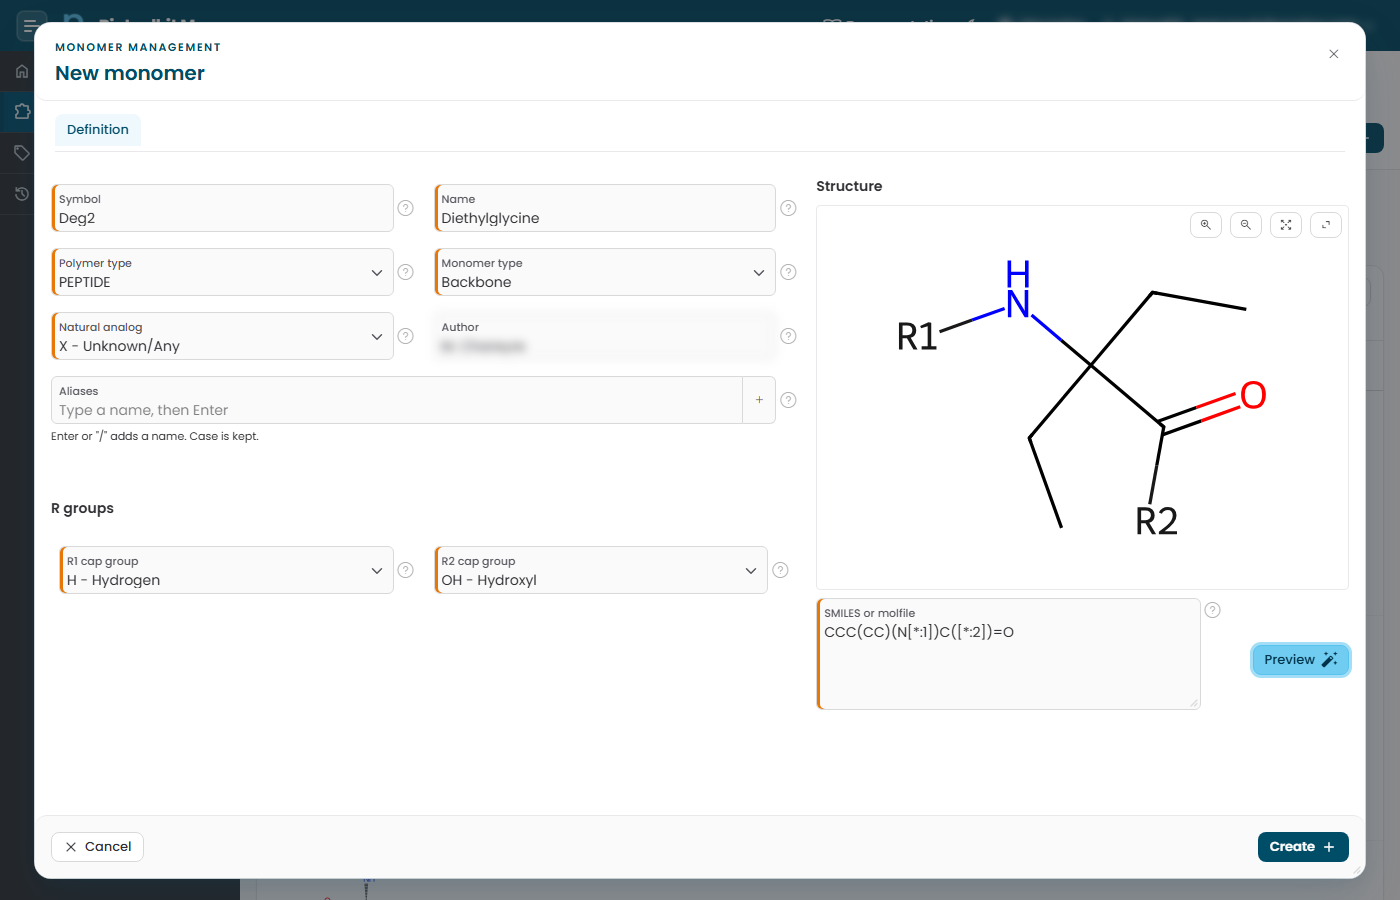
Check three things on the drawing:
- the skeleton is the one you meant;
- the R labels sit on the right atoms;
- the number of R labels matches the number of
[*:n]in your SMILES.
If the message reads Invalid SMILES. Could not generate structure., the string is not parseable. Common causes are an unbalanced bracket or parenthesis, a lowercase aromatic atom outside a ring, or an attachment point written without a number. Fix the text and click Preview again.
The puzzle button above the drawing shows the structure with its cap groups applied, as they are chosen in the R groups section. It follows your choices before you save, on a new monomer as on an existing one.
Starting from a molfile
You can paste a molfile (V2000 or V3000, from Ketcher, ChemDraw, Marvin or a file) into SMILES or molfile instead of a SMILES:
- Paste it. The molfile stays in the field as pasted.
- Click Preview. The application reads it and replaces it with its CXSMILES, then draws it. R-group atoms (
R1,R2, a genericR) and Ketcher attachment points become the[*:n]attachment points. - If the molfile draws stereocentres without the chiral flag, a question asks how to register it: Racemic (&1), the MDL standard reading, or As drawn (absolute), for a file from a program that does not set the flag for a single enantiomer. Ketcher sets the flag, so a Ketcher molfile does not ask. See Enhanced Stereochemistry.
Create and Save changes refuse a molfile that has not been previewed, with Structure has changed. Please click Preview before saving. under the field. Nothing is saved. A molfile with enhanced stereo on bonds (a STEBABS, STEBRAC or STEBREL collection) is refused at Preview, because the library cannot store it, and stays in the field: Enhanced stereochemistry on bonds (MDLV30/STEBRAC1) is not supported: the molfile would be registered as the isomer drawn. Register each isomer as its own monomer, or remove the collection.
Step 5: Set the cap groups
For each R-group, choose the cap group: the small group that completes the valence when this attachment point is not used in a chain. For a standard peptide monomer:
| R-group | Cap group | Why |
|---|---|---|
| R1 | H - Hydrogen |
A free N-terminus is an amine, NH2. |
| R2 | OH - Hydroxyl |
A free C-terminus is a carboxylic acid, COOH. |
Every selector opens on H - Hydrogen, so set R2 cap group to OH - Hydroxyl for Deg. The vocabulary also offers NH2 - Amino, Azide, Ethynyl, Ethenyl - Vinyl and the halogens Br, Cl and I. See Attachment Points and CAP Groups.
Each click on Preview rebuilds the cap group selectors and puts them back to H. Set the caps after your last preview, just before you save.
Step 6: Create
Click Create. The service validates the structure, computes its canonical form, generates the drawing, and the dialog closes. The new monomer appears in the Monomers report with its structure, symbol, types and R groups, and the Private chip of the Scope filter now counts it.
These messages can stop the creation here:
- Structure has changed. Please click Preview before saving. You edited the SMILES after the last preview. Preview again, then create.
- "Diethylglycine" is already the name of Deg2 in your organization. A name is used only once per polymer type., under the Name field. Another of your monomers has this full name. Change the name, or open that monomer.
- Monomer with this structure is already registered: meG. If you expected a distinct structure, verify your input carries full stereochemistry … The same molecule already exists, here as the public monomer
meG. Use that symbol instead, or add your symbol to it as an alias: see How to Manage Monomer Aliases and Canonical SMILES and Duplicate Detection. - Monomer with this symbol is already registered: …, or Cannot modify protected public monomer '…' (public seed data). The symbol is taken, by one of your monomers or by a public one. Pick another symbol.
- "Deg" is already an alias of monomer Et2G. The symbol is already a name of another monomer, as an alias. A name points to one monomer per polymer type: pick another symbol, or use that monomer.
- Axial chirality is not supported (the stereo on atom 4, in a spiro ring system) … The structure is a hindered biaryl, an allene or a 2,6-disubstituted spiro[3.3]heptane drawn with its axial stereo, which the canonical key cannot keep. Register it without that stereo and tell the forms apart by their name; see Enhanced Stereochemistry.
- Registering 'Deg' corresponds to 1 deleted monomer (2504). Nothing was written. A monomer with this symbol or this structure was deleted earlier. Ask an Administrator to revive it from the Version history rather than registering a second one. See Versioning and Deletion.
Editing a monomer: correcting a cap group
Suppose you registered N-methyl-norleucine as MeNle, previewed it, and clicked Create without going back to the cap groups. R2 was saved as H, so the free residue is an aldehyde and every peptide ending on it gets a wrong molecular weight.
- On the Monomers page, find
MeNle, with the search box or the chips, and click its symbol or the pencil icon in the Actions column. The Monomer management dialog opens on the monomer, with its symbol, name, polymer type, version and scope in the title. Symbol and Polymer type are locked; the other fields stay editable, and the R groups selectors show the saved values. - Set R2 cap group to
OH - Hydroxyl. Do not click Preview, unless you also changed the structure; if you do, set the caps again afterwards. - Click Save changes.
The cap groups count as a material change, so the monomer's version number goes from 1 to 2 and the previous state is archived. The Audit tab of the dialog lists the versions of this monomer, and an Administrator can see the before and after in the Version history, with What changed reading R-groups; see How to Review a Monomer's Version History. Changing only the Name or the Author does not create a version, and neither does adding or removing an alias. On an existing monomer an alias is saved as soon as it is added, without Save changes; see How to Manage Monomer Aliases.
The SMILES shown when you reopen a monomer is the normalised form computed by the service, not the text you typed. It is chemically the same molecule. The buttons under the structure copy it as SMILES, CXSMILES or molfile.
Deleting a monomer
Deleting is reserved to Administrators: the bin icon in the Actions column of the Monomers report, and the Delete button of the dialog, are active only for them and only on a private monomer. Click it and confirm in the Delete monomer dialog, which reads Delete this monomer? It will disappear from the catalogue; a Monomer Administrator can restore it from the Version Log. The monomer disappears from the library, and its full state is archived so that it can be revived later. If the monomer still belongs to a monomer set, the deletion is refused with the list of sets to remove it from first. See Versioning and Deletion.
Next steps
- Register many monomers at once: How to Import Monomers in Bulk.