This guide covers importing a peptide dataset from a HELM file (CSV/Excel) or an SDF, registering its custom monomers, configuring peptide-specific options, reading the dataset table, and downloading the dataset.
The Peptide dataset type is only available when the peptide plugin is enabled for your organization. If you do not see it, contact your administrator.
Step 1: Open the New Dataset wizard
From the Home page, click the Create Dataset card (or New → Create Dataset in the left navigation). The New Dataset wizard opens with three steps: Dataset Information, Properties Configuration, and Import Status.
Step 2: Choose the project and the dataset file
Choose the target Project, then drop your file on Dataset File (or click it to browse). Ideation reads the file, sets the Dataset Type for you and tells you when it switched it, with "Detected a Peptide dataset. You can change the type above."

| File you drop | Dataset Type selected |
|---|---|
| CSV with a HELM column | Peptide |
| CSV without a HELM column | Small Molecule |
Excel (.xlsx) |
Peptide |
| SDF whose first molecule carries monomer templates | Peptide |
| Any other SDF | Small Molecule |
Compressed file (.csv.gz, .sdf.gz, .sd.gz) |
Not read: the selected type is kept |
Check that Dataset Type reads Peptide before you continue, and select it yourself if the guess is wrong or your file is compressed. If the type does not accept the file's format (an Excel file with Small Molecule, for example), Next stops with "This file format is not supported for Small Molecule datasets."
Accepted file formats for peptides:
| Format | Notes |
|---|---|
CSV / Excel (.csv, .csv.gz, .xlsx) |
Tabular files with an identifier column and a HELM column; every other column becomes a property. HELM references monomers by symbol only, so each monomer must be registered in Biotoolkit Monomers before the import (Step 3). |
SDF (.sdf, .sd, .sdf.gz, .sd.gz) |
A V3000 SDF that carries the peptides and a monomer template for each monomer. The embedded templates are registered in Biotoolkit Monomers during the import, so nothing needs registering beforehand. |
With the Dataset Type set to Peptide, use the Download templates links (CSV / Excel) under the file picker to start from a correctly formatted file. The template's columns are ID, HELM and three example property columns, so a minimal row looks like PEP001,PEPTIDE1{A.G.K.L}$$$$. For branches, bridges, CHEM monomers and the HELM features Ideation does not support, see HELM Rules for Peptide Import.
Step 3: Register your custom monomers (CSV/Excel imports only)
HELM names each monomer by its symbol only. For a CSV or Excel file, Ideation reads the structure of each monomer from your organization's monomer library in Biotoolkit Monomers, so register your custom monomers there before the import. Under the file, the notice Monomers must be registered in Biotoolkit Monomers links to the app; point to the notice for a short explanation.
- Natural amino acids and the public monomers are usually in the library already. Only the monomers your organization invented need registering.
- To register many monomers at once, import a monomer file in Biotoolkit Monomers itself.
- A peptide that uses a monomer missing from the library is still imported: it keeps its HELM sequence but gets no structure, and the import issues list the missing monomer. Register it in Biotoolkit Monomers, then run Sync monomers. See How to Resolve Missing Monomers.
A V3000 SDF embeds a monomer template for every monomer. During the import, Ideation registers the templates of the monomers your peptides use in Biotoolkit Monomers, which is why the notice does not appear for SDF files.
Step 4: Configure peptide-specific options
Click Next. On the Properties Configuration step, a Peptide Specific Configuration section appears:
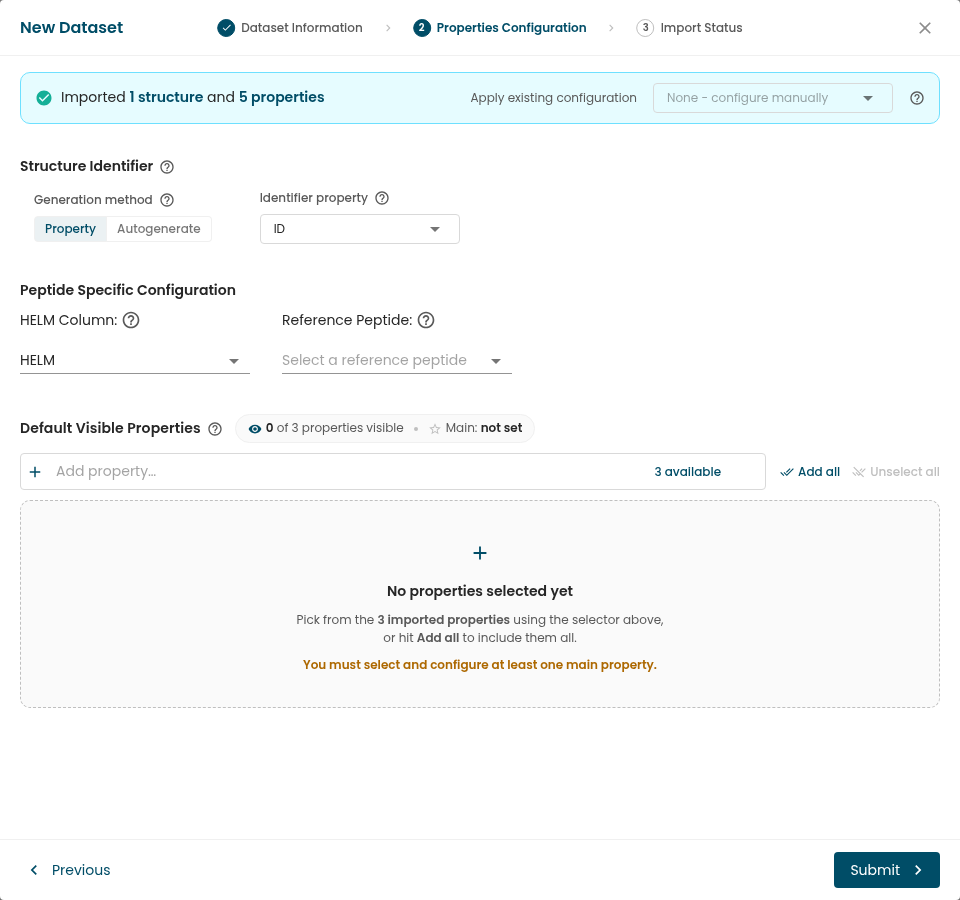
- HELM Column — the column that holds the HELM sequence notation (tabular imports only; auto-detected when named
HELM). - Reference Peptide — the peptide used by default as the basis for SAR slides created from this dataset. You can change it later per report.
Select and configure at least one main property, then submit.
Step 5: Watch the import
The final Import Status step shows the peptide import timeline:
- CSV or Excel file: Initializing → Importing Peptides → Ready
- SDF: Initializing → Registering Monomers → Importing Peptides → Ready

- Registering Monomers registers the monomer templates of a V3000 SDF in Biotoolkit Monomers.
- Importing Peptides converts the structures and computes their properties. A progress bar covers the whole file and names the current step, for example Peptides 1–500 of 1,223 · step 3/4: converting the structures.
When the dataset reaches Ready, it opens on the dataset page. Its alignments are computed next, in the background: a Center-star alignment and a PepSeA alignment appear on the dataset's Alignments tab and complete on their own, so you can build a SAR Slide without providing an alignment file.
An SDF import never changes a public monomer: its registered structure is used. A monomer of your organization's library is different: when the SDF embeds a template with the same symbol but another structure, the import updates that monomer, and Biotoolkit Monomers keeps its previous version. The change applies to every dataset that uses the monomer, so check the templates of an SDF before you import it.
Biotoolkit Monomers checks the structure of each new monomer. When the same monomer already exists under another name, your symbol is added to it as an alias instead of registering a duplicate. For example, if your file names a monomer Zzz2 and the service already holds the same structure as Zzz9, Zzz2 becomes an alias of Zzz9. Your peptides keep the symbol you wrote. The alias applies to one polymer type (PEPTIDE or CHEM), because a symbol can name a residue and a different CHEM monomer. You can see it with the monomer in Biotoolkit Monomers. When your symbol cannot become an alias, for example because another monomer already uses that name, the monomer is reported in the import issues dialog. See How to Resolve Missing Monomers.
An alignment that cannot run ends as a Failed row on the Alignments tab, and the import still succeeds. Center-star does not run on datasets with no HELM sequences or above 5,000 peptides; PepSeA does not run on datasets with too many distinct non-natural monomers. See How to Align Peptides.
If some peptides reference monomers that could not be resolved, they are imported with their HELM sequence preserved but without full structure or properties, and the dataset is flagged with an amber import issues warning that lists them under Peptides imported with missing data. See How to Resolve Missing Monomers.
Step 6: Read the dataset table
The Peptides tab of the dataset page lists one row per peptide, and the table footer counts them (Peptides: N). Besides your own properties, a peptide dataset carries these common columns:
| Column | Shown by default | What it holds |
|---|---|---|
| ID | yes | The primary identifier. |
| Backbone Sequence | yes | The peptide drawn as its sequence of monomers. |
| Num Monomers | yes | How many monomers the HELM declares. |
| Num Atoms | yes | The number of heavy atoms (every atom except hydrogen) of the peptide. |
| Monoisotopic Molecular Weight | no | The exact mass, computed with the most abundant isotope of each element. |
| Average Molecular Weight | yes | The molecular weight computed with average atomic masses. |
| InChI String | no | The InChI of the whole peptide. |
| InChI Key | no | The InChIKey of the whole peptide. |
| Canonical SMILES | no | The canonical SMILES of the whole peptide. |
| HELM | yes | The peptide's HELM notation. When the conversion succeeds and no monomer had to be inlined, its chains are renumbered in the order your file writes them, so a chain number can differ from your file — see HELM Rules for Peptide Import. Otherwise, the HELM is stored as written. |
| Missing Monomers | when needed | The unresolved symbols of a peptide imported without its structure; the column exists only while something is unresolved. |
Ideation computes the atom count, the weights, the InChI, the InChI Key and the canonical SMILES from the peptide's structure. They are empty for a peptide whose structure could not be converted.
To show a hidden column, open Table settings on the Peptides tab, then add the column to Displayed Columns with its add button (Add columns to display). The InChI, InChI Key and canonical SMILES start hidden because they are long identifiers you copy rather than read.

On a dataset imported before the 4 November 2026 release, Average Molecular Weight and InChI String are empty, and Num Atoms also counts the hydrogen atoms at the ends of the peptide. Run Sync monomers to fill and correct them (see How to Resolve Missing Monomers).
The row menu (the list icon in the Actions column of a row) offers Create SAR Slide with that peptide as reference. It also copies the peptide to the clipboard: Copy HELM, Copy CTAB v2000, Copy CTAB v3000, Copy Canonical SMILES, Copy InChI String and Copy InChIKey. A copy action is greyed out when the peptide has no such value, for example when its conversion failed at import. Hover it to read why.
Formula columns work on a peptide dataset as on any other: add one with New Formula Column on the Endpoints tab of the dataset settings drawer. Besides your own columns, a formula can use ID, Monoisotopic Molecular Weight, Average Molecular Weight, Num Atoms and Num Monomers. For example, a formula that returns the average molecular weight lets you use the weight as an endpoint in a SAR Report. Sync monomers recomputes the formula columns that use these values. If it cannot recompute one, the column stays empty and the import issues dialog tells you why.
Download a peptide dataset
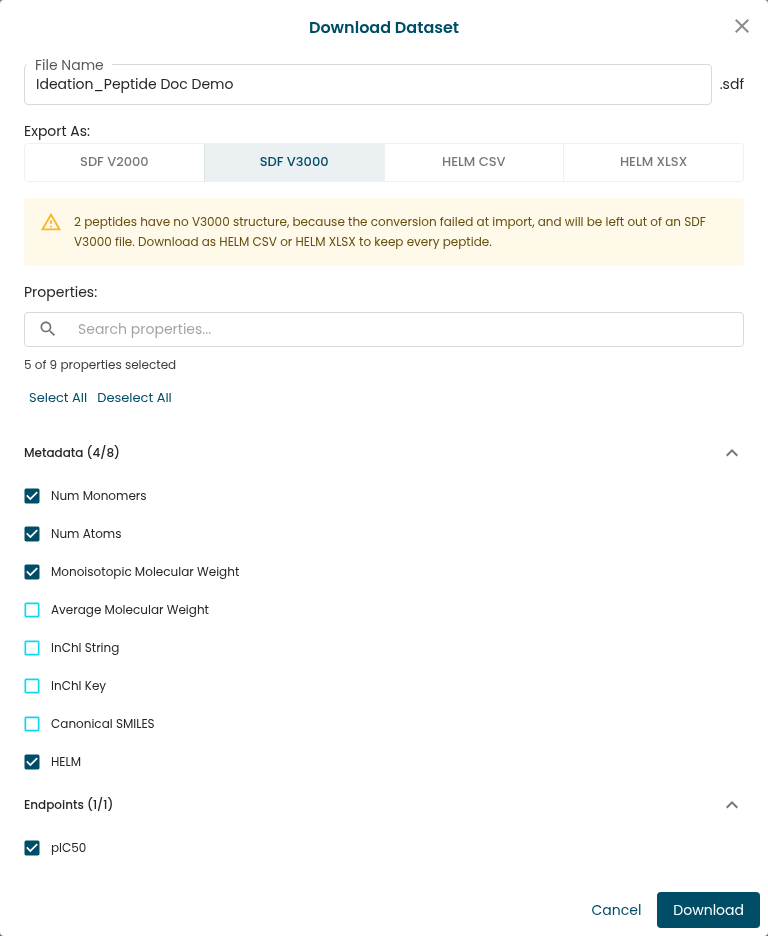
Click the download icon (Download dataset) in the dataset title bar. It is available once the dataset is Ready. In the Download Dataset dialog:
-
Change the File Name if you want.
-
Under Export As:, choose the format:
Format What the file holds SDF V2000 One record per peptide, with its full-atom structure. SDF V3000 One record per peptide, with a structure that keeps each monomer as a monomer template. This is the format selected first. HELM CSV One row per peptide, with its HELM. HELM XLSX The same as HELM CSV, as an Excel file. -
For HELM CSV, choose the separator under CSV separator:. Comma (EN) writes decimals with a point (1.5), for English-language Excel and most tools. Semicolon (FR) writes decimals with a comma (1,5), for French-language Excel. The dialog starts on your last choice, or on your browser's language.
-
If the table is filtered, choose under Structures: All to download every peptide, or Filtered to download only the peptides the filters keep.
-
Under Properties:, tick the columns to write. Metadata lists the values Ideation computes (Num Monomers, Num Atoms, the two molecular weights, InChI String, InChI Key, Canonical SMILES, and HELM for an SDF). The ones shown in the table are ticked. Endpoints lists your own columns, all ticked.
-
Click Download.
A peptide whose conversion failed at import has no structure. An SDF V3000 file leaves it out, and an SDF V2000 file writes it with an empty structure. The dialog tells you how many peptides this concerns. Choose HELM CSV or HELM XLSX to keep every peptide.
A HELM CSV, HELM XLSX or SDF V3000 file downloaded from Ideation imports back as a peptide dataset, without any change. Ideation computes the Metadata columns again, so it does not import them as properties. The Properties Configuration step lists the columns it leaves out ("Columns recomputed by Ideation are not offered as properties: …"), and so does the import issues dialog of the new dataset. Only the exact column names Ideation writes are left out: a column of your own named InChI, for example, still imports as a property.